gcorr software validation
These recorded simulation results assess selected behavior on ten fixed datasets from the shared design. They do not revalidate the statistical method or cover every setting. The use snippets show the current package interfaces. This report step summarizes retained artifacts and does not rerun analyses. The gcorr guide describes current inputs and options.
Use
genome <- gcorr::gcorr(statistics, ld_scores = ld_scores, blocks = blocks,
                       method = "ldsc")
partition <- gcorr::gcorr(statistics,
  annotation_ld_scores = annotation_ld_scores,
  weight_ld_scores = ld_scores, annotation_sums = annotation_sums,
  blocks = blocks, reference_sample_size = reference_sample_size,
  method = "ldsc")
Results
Trait A heritability and A–B genetic correlation are shown below. Other traits, regions and uncertainty availability remain in the compact result tables.
| Method | h2 estimate | h2 truth | h2 bias | rg estimate | rg truth | rg bias |
|---|---|---|---|---|---|---|
| LDSC | 0.263 | 0.300 | -0.037 | 0.551 | 0.564 | -0.013 |
| GNOVA | 0.259 | 0.300 | -0.041 | 0.553 | 0.564 | -0.012 |
| SumHer | 0.263 | 0.300 | -0.037 | 0.551 | 0.564 | -0.013 |
| Annotation LDSC | 0.347 | 0.300 | 0.047 | 0.547 | 0.564 | -0.018 |
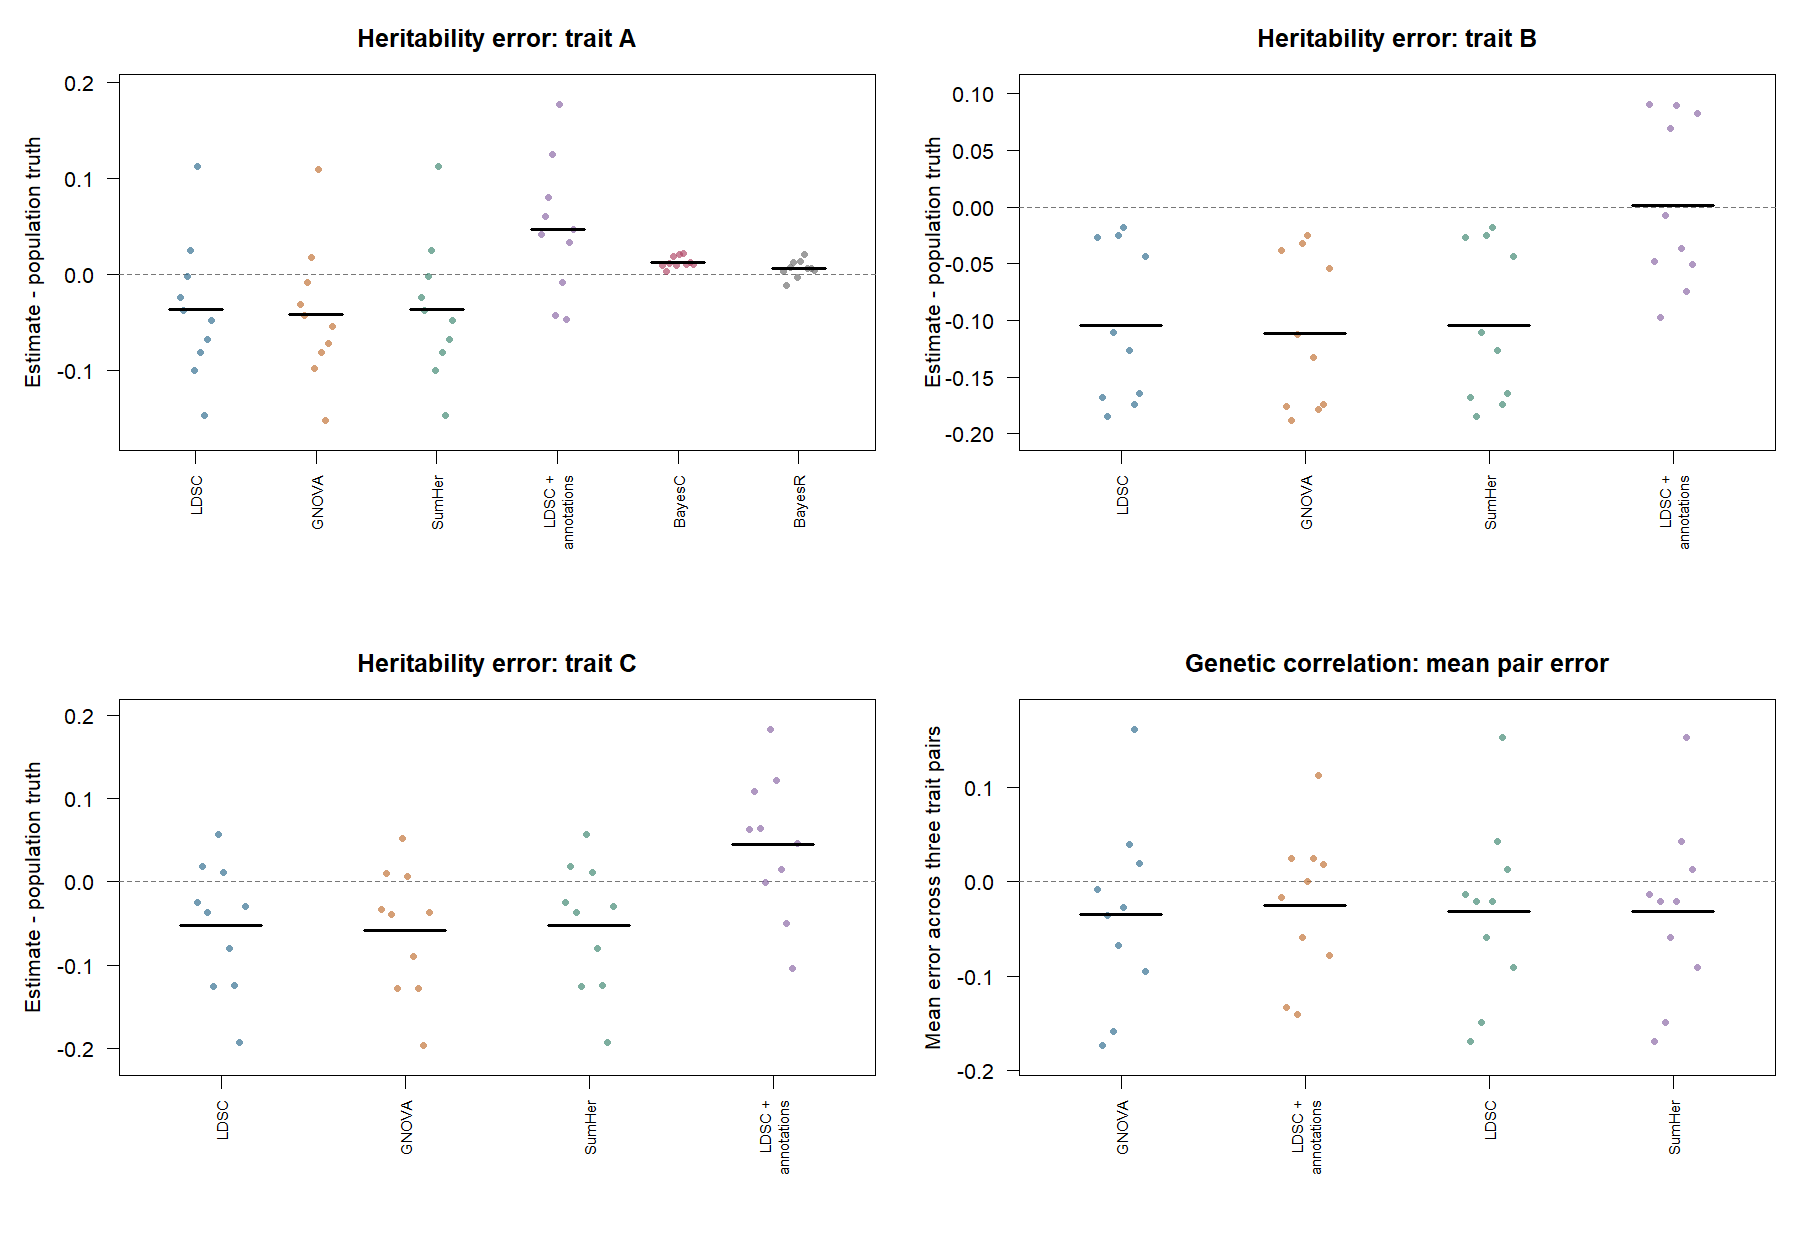
These methods have different assumptions and uncertainty constructions. Ten replicates do not establish interval coverage or rank methods. Estimates that were unavailable or outside the correlation range are retained in the results.
Seed-level estimates · Accuracy summary · Uncertainty availability